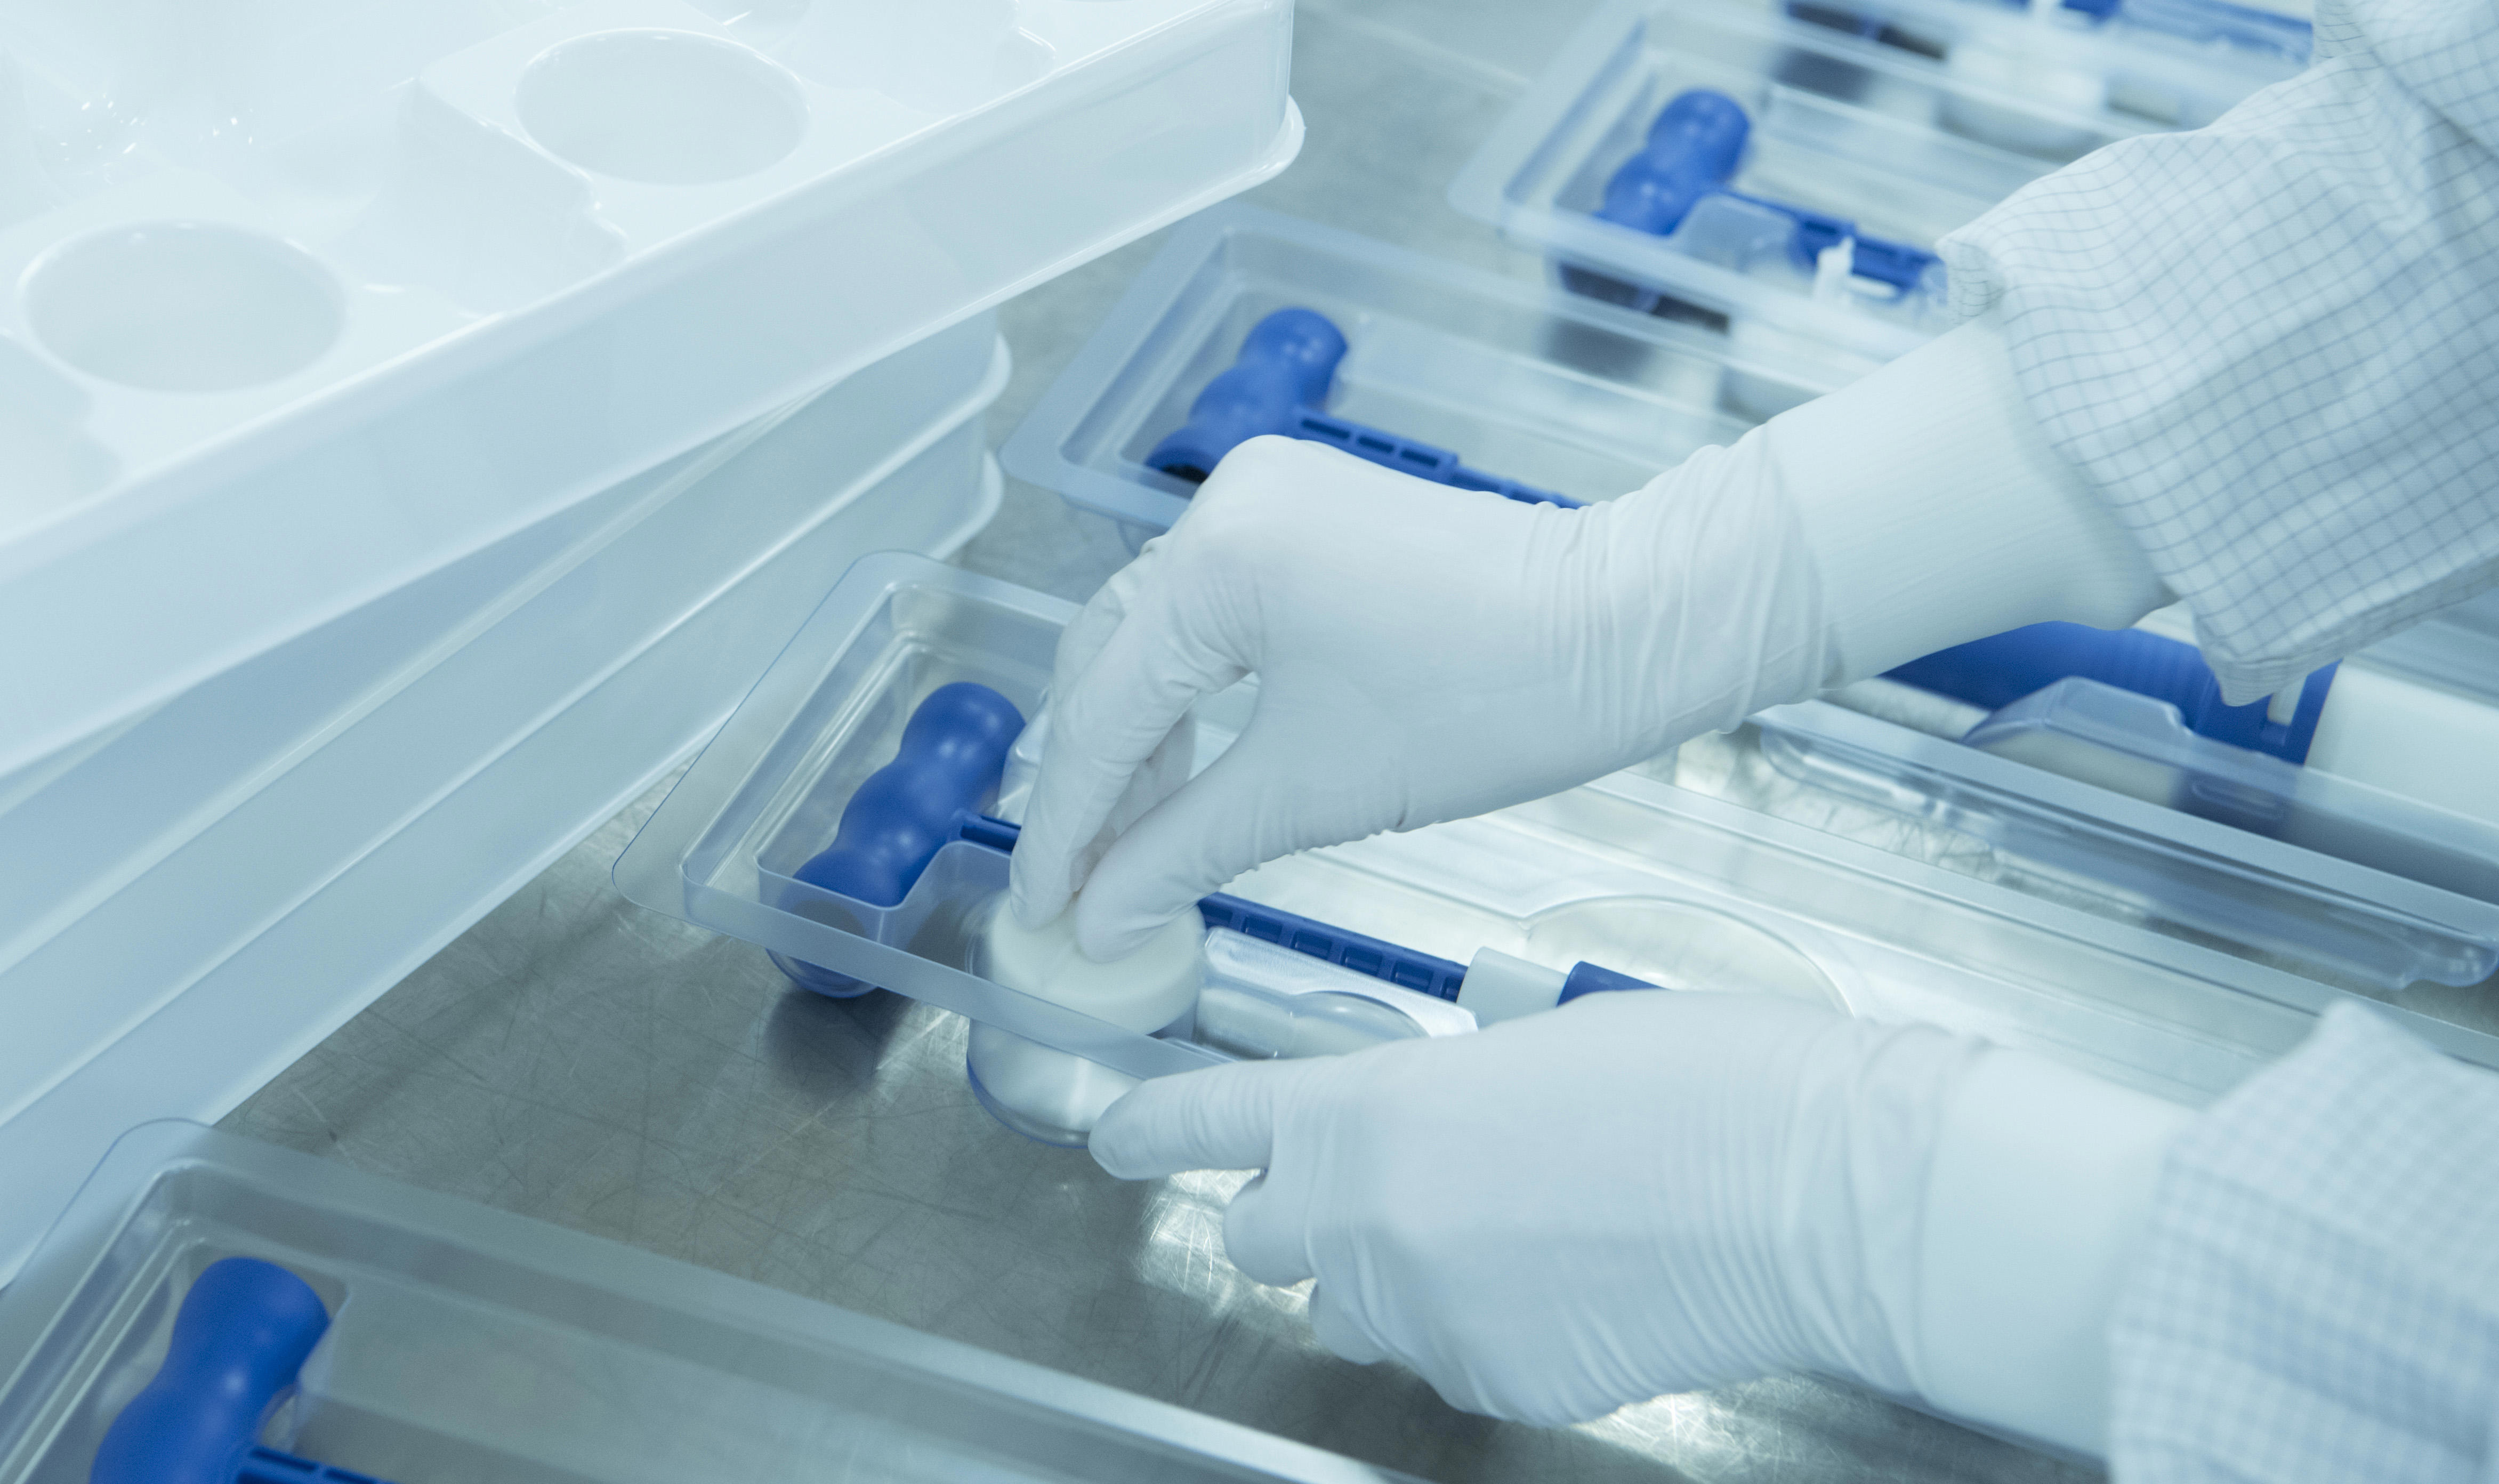
 For manufacturers of medical devices, packaging validation is often perceived as a complex regulatory challenge. While significant attention is typically paid to product development, risk management, and sterilisation processes, packaging is frequently neglected and only addressed shortly before market launch. This can lead to unexpected delays, costly redesigns, and difficulties during regulatory reviews. ISO 11607, the international standard for packaging of sterilised medical devices, provides a structured framework. However, many organisations struggle to translate the standard’s requirements into practical actions.
A common misconception is that packaging validation is a standalone activity. Packaging must be considered throughout the entire medical device lifecycle. Decisions made during product development, such as material selection, geometry, or sterilisation method, are all interconnected and have a direct impact on the packaging concept. A packaging material that performs well in product protection may be incompatible with a chosen sterilisation method. Considering these interactions early in development helps avoid costly changes later in the project.
Selecting an appropriate sterile barrier system is one of the first critical steps in packaging development. ISO 11607 does not prescribe a specific packaging design but requires manufacturers to demonstrate that the selected system is suitable for its intended purpose. Common solutions include thermoformed blisters, soft blisters, and pouch systems. Thermoformed blisters provide excellent protection for fragile, sharp, or geometrically complex devices because they maintain a defined product position, while pouches offer flexibility and cost efficiency and are often suitable for less sensitive products. The optimal solution depends on product characteristics, sterilisation compatibility, transport conditions, usability requirements, and risk management considerations.
Beyond physical protection, packaging materials must fulfil additional requirements. They must provide an effective microbial barrier, remain compatible with sterilisation processes, and maintain their properties throughout the intended shelf life. Biocompatibility considerations are equally important, particularly for materials in direct contact with the medical device. Furthermore, the packaging should enable intuitive and aseptic presentation of the device at the point of use.
Understanding the structure of ISO 11607 is essential for navigating its requirements. Part one focuses on the materials, while part two addresses validation process and provides the framework for the validation of forming, sealing, and assembly processes. Together, both parts ensure that manufacturers validate not only the package itself but also the process used to produce it.
One of the most valuable concepts within ISO 11607 is the risk-based worst-case approach. Packaging systems with similar characteristics can often be grouped into families, allowing representative configurations to be validated. Identifying the worst-case scenario requires a documented rationale that considers factors such as package dimensions, material combinations, and tooling. This approach increases efficiency while maintaining a high level of confidence in product safety.
Packaging validation follows the sequence of Installation Qualification (IQ), Operational Qualification (OQ), and Performance Qualification (PQ). IQ confirms that equipment and environmental conditions meet predefined requirements, OQ demonstrates that the process performs reliably across defined limits, and PQ verifies that routine production delivers compliant sterile barrier systems under standard operating parameters.
Performance testing plays a central role throughout validation. Depending on the packaging design, common methods include seal strength testing, dye penetration testing, visual inspections, and peel assessments. Statistical justification of sample sizes and acceptance criteria is essential to ensure valid results.
However, packaging validation does not end once the sealing process has been validated. The sterile barrier system stability must also be demonstrated after the processes of sterilisation, transportation, and long-term storage. Worst-case sterilisation studies, transport simulation, and ageing studies provide evidence to attest to package integrity throughout the product lifecycle.
Navigating ISO 11607 becomes significantly easier when packaging is viewed as an integral part of product development. By combining early planning, risk-based decision making, process validation and lifecycle thinking, manufacturers establish sterile barrier packaging systems that protect products, maintain sterility, and ensure patient safety. This approach turns the apparent jungle of ISO 11607 into a clear and manageable pathway towards compliance and successful market access.
For more information visit: www.eurofins.com/en/biopharma-services/medical-device-services/